Health Canada Medical Device Registration & Approval (CMDR)
Last reviewed:
Plan Canadian medical device access with the correct Class I–IV pathway, Medical Device Licence and establishment-licensing assessment. Pure Global supports Health Canada submissions, quality-system readiness, labeling and lifecycle compliance.
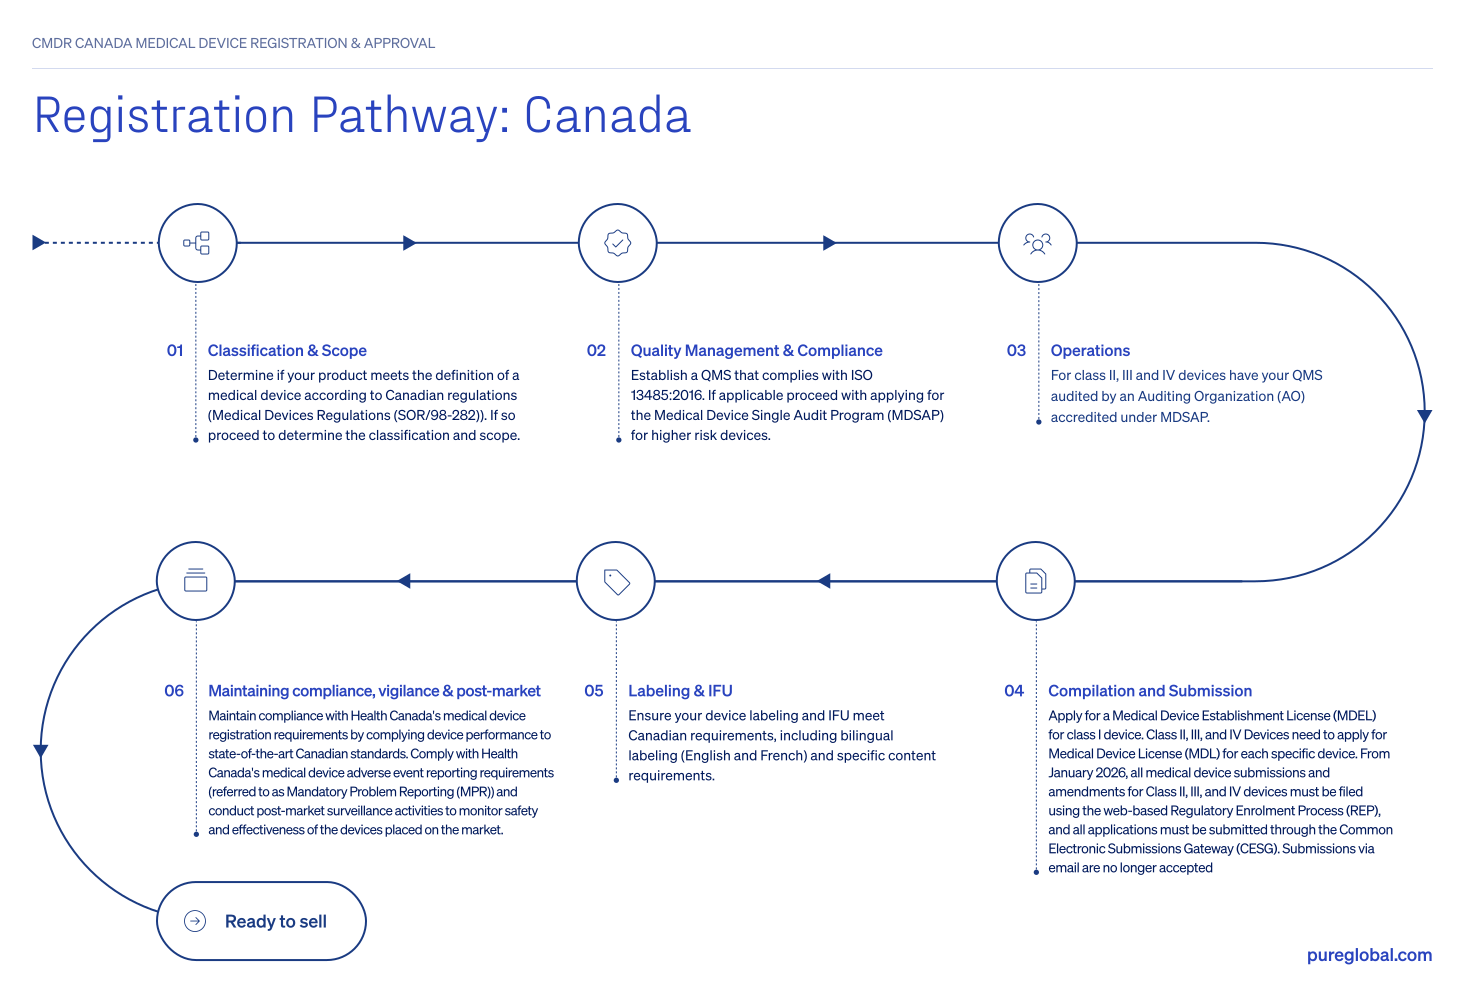
Canada Medical Device Registration Process (Health Canada)
Canada medical device registration: MDL and MDEL are different
Health Canada regulates devices under the Medical Devices Regulations. Class I products generally do not need a product-specific Medical Device Licence (MDL); Class II, III and IV products do. Establishment licensing depends on the activities performed by manufacturers, importers and distributors, with specific exemptions.
An MDEL is not approval of each product. An MDL does not automatically license every importer or distributor in the supply chain. Define the legal manufacturer, device class, import route and commercial roles before preparing applications.
Classification, licence and quality evidence
| Device or activity | Main requirement to assess | Evidence and planning point |
|---|---|---|
| Class I product | No MDL; assess MDEL obligations for the relevant establishment | Classification rationale, labels and operating procedures still matter |
| Class II product | Manufacturer’s MDL | Required attestations and applicable quality certificate |
| Class III or IV product | Manufacturer’s MDL with class-appropriate supporting information | Safety/effectiveness evidence and structured technical documentation |
| Import/distribution | MDEL unless an exemption applies | Activities, device classes, sites and documented procedures |
Apply the Canadian classification rules, including the separate IVD rules, rather than importing a US or EU class. Class II–IV manufacturers generally need an ISO 13485 certificate issued through the Medical Device Single Audit Program (MDSAP) for the licensed scope. Confirm manufacturer/site identity and device coverage before using a certificate in the application.
Submission planning: REP and CESG
Health Canada’s mandatory REP notice took effect April 1, 2026 for the covered medical-device regulatory transactions. REP captures enrolment and application information; the Common Electronic Submissions Gateway (CESG) transmits the transaction. Do not describe every Canadian medical-device process, including MDEL, as the same product-licence submission workflow.
Establish the required company/dossier identifiers and gateway access early. Reconcile device names, identifiers, intended use, class, manufacturer, quality certificate and application category. Prepare the applicable structure and attachments, submit, retain acknowledgments and map questions to controlled responses. A successful electronic transmission is not a licensing decision.
Canadian labeling and local responsibilities
Sections 21–23 govern labeling. The language rule distinguishes products sold to the general public, for which specified information must be in both English and French, from other devices where English or French may initially suffice subject to the applicable obligations. Assess provincial and commercial requirements separately. Do not claim one identical bilingual rule for every professional-use device.
Canada does not impose one universal EU-style local Authorized Representative on every foreign manufacturer. A foreign manufacturer can hold an MDL, while importers/distributors and their MDEL duties must be assessed. Contracts should define complaint escalation, mandatory reporting, distribution records, recall cooperation, record access and who communicates with Health Canada.
Renewal, fees and postmarket compliance
Active MDEL holders generally submit annual licence review before April 1 and pay under the invoice terms; the application deadline is not automatically the fee-payment deadline. Licensed Class II–IV manufacturers have an annual renewal notification before November 1, with applicable right-to-sell fees handled under the current process. Maintain current MDSAP evidence, licence amendments and supply-chain permissions.
Government application fees, annual fees, audit costs and consulting costs are separate. Review standards depend on application type and exclude some applicant work; they are not a guaranteed total launch time. Budget for enrolment, testing, MDSAP readiness, dossier preparation, review questions and labeling changes.
After licensing, assess design, intended-use, site and quality-certificate changes before implementation. Maintain incident reporting, recall and safety-information procedures. Use the MDL/MDEL guide for the activity matrix and our pricing information to define the service scope. Pure Global can support the pathway and application while the manufacturer and licensed establishments retain their responsibilities.
 Regulatory Update · Sep 10, 2025Health Canada eSTAR Pilot 2025: Streamlining Class III & IV SubmissionsBlog · Aug 24, 2026Medical Device Registration Verification 2026Blog · Aug 17, 2026Coronary Stents: Eight Highest-Tier FilesBlog · Aug 10, 2026Surgical Robots: Eight Markets, Eight Regulatory FilesBlog · Aug 3, 2026ISO 10993-1:2025: One Core, Seven Regulatory SystemsVideo · Oct 16, 2026Health Canada: MDEL and MDL ExplainedVideo · Oct 16, 2026ISO 10993-1:2025: What ChangedVideo · Jul 28, 2025How to Register a Medical Device in CanadaRegulatory Update · Oct 17, 2026FDA Toxicology 510(k) ExemptionsRegulatory Update · Oct 11, 2026ANMAT Foreign Plant GMP Certificates
Tailored Support for Canadian Market Entry
We offer comprehensive support to streamline your market access in Canada:

Classifying and grouping your products to define the regulatory pathway.
Assisting with the compilation of the technical documentation for MDL or MDEL applications.
Auditing your QMS to Health Canada's MDSAP requirements.
Frequently asked questions
An MDL licenses a Class II, III or IV device for its manufacturer. An MDEL concerns an establishment’s permitted activities and operating controls. Device class, supply-chain role and exemptions determine which licences are needed.
Health Canada’s notice specifies April 1, 2026. Covered transactions use REP and CESG; do not extend that statement to every establishment-licensing or special-access process without checking the applicable instructions.
The federal language rule distinguishes general-public devices from others. Specified public-use information must be in English and French; professional-use requirements differ, and provincial or other applicable requirements should also be assessed.
No. Foreign approvals may support an eligible evidence or review strategy, but they do not automatically replace the Canadian licence. Confirm the device class, applicable route, required evidence and Canadian labeling requirements.
Single Process,
Multiple Markets
When you partner with Pure Global, a single registration process opens doors to multiple countries. Our global subsidiaries make this streamlined path possible.








Let's Talk,
Anywhere You Are.
Whether looking for more information or ready to partner with us, we're here to guide you through every step of the regulatory process.
Contact us