MDD Hong Kong Medical Device Registration
Last reviewed:
Plan Hong Kong MDACS listing with the applicable reference evidence, Local Responsible Person, technical file and postmarket controls. Assess current procurement requirements separately from voluntary listing.
Video guide
Hong Kong MDACS: Voluntary Listing
Explore Hong Kong's MDACS listing framework, the CMPR and the changes covered in this video.
Open video pageTranscript (English)
Is Hong Kong listing voluntary? For now, yes. But a new regulator, the CMPR, is being set up this year, and device legislation is planned. Today, the Medical Device Division runs MDACS. A Local Responsible Person in Hong Kong applies online. Class II to IV devices and Class B to D IVDs can be listed, using a conformity assessment certificate or an approval from eight recognised jurisdictions. The listing itself is free, normally vetted within 12 weeks, and valid for 5 years. But renewal has a strict window: 12 weeks to one year before expiry. Miss it, and you start again. Change is coming. The government plans a law requiring registration of all devices supplied in Hong Kong, unless exempted. On October 12, it named a Controller for Medical Products Regulation. Start with three things. One: appoint a Local Responsible Person in Hong Kong. Two: gather a recognised approval or a CAB certificate. Three: put the renewal window in your calendar. List now, prepare for registration. Pure Global covers registration, in-country representation, quality systems, audits and cybersecurity, in more than 30 markets. Visit pureglobal.com.
Navigating Regulatory Requirements in Hong Kong
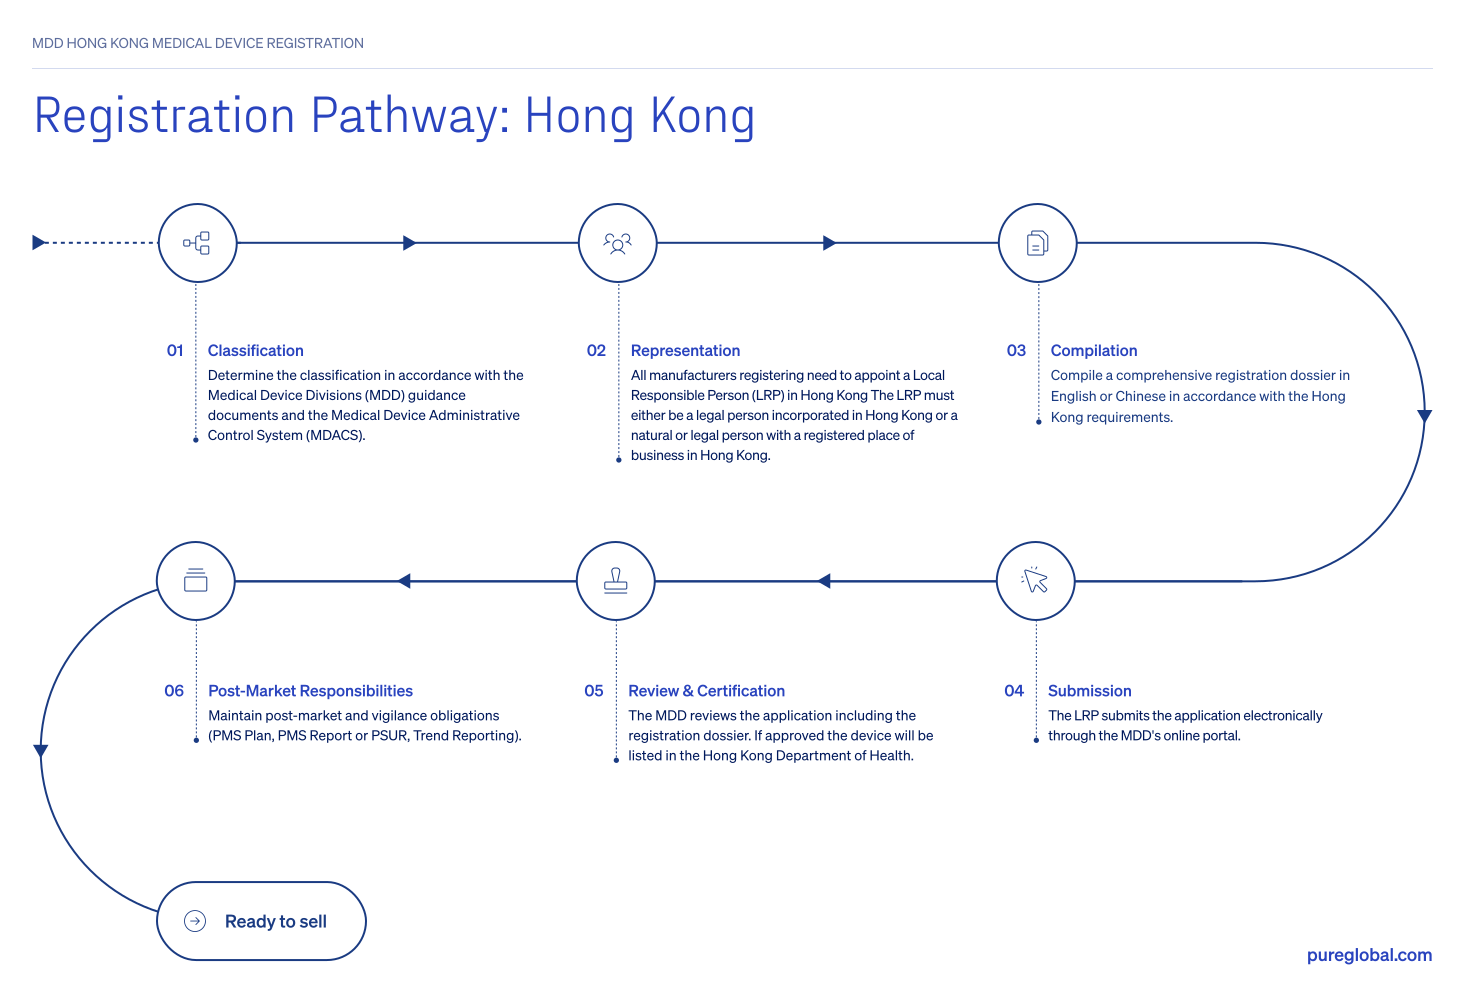
Medical devices and IVDs in Hong Kong are overseen by the Medical Device Division (MDD). MDACS is an administrative listing and monitoring framework supported by Guidance Notes and Technical References; other applicable laws and procurement requirements still need assessment.
Key documents include:
Hong Kong’s medical device regulatory framework is established under the Medical Device Administrative Control System (MDACS), which includes a device listing system (via the Medical Device Information System (MDIS)) and an adverse event reporting system. MDACS listing is voluntary under the administrative system, while specific public procurement requirements can make listing important for a particular tender. Do not equate that with one universal requirement for every hospital purchase.
All devices listed by manufacturers with no entity in Hong Kong need a Local Responsible Person (LRP). Only the LRP can submit the listing application, and the LRP has defined pre- and post-market obligations.
Hong Kong Medical Device and IVD Classification
Hong Kong classifies medical devices and IVDs in line with IMDRF principles. Medical devices are classified as Class I, II, III, and IV; IVDs are classified as Class A, B, C, and D.
Hong Kong (MDD) medical device regulatory pathways
MDD has established MDACS listing pathways for Class II/III/IV general medical devices and Class B/C/D IVDs based on existing reference country approval and market history. Class I devices are not listed. MDACS listing requires a Conformity Assessment Certificate from an MDACS-accredited conformity assessment body (CAB). CAB review is not required for devices with approval in one of Hong Kong’s approved reference markets: Australia, Canada, the European Union, Japan, the United States, Mainland China, South Korea, or Singapore.
Standard MDACS listing route (with and without reference market approval)
Process: the LRP applies for inclusion into the List of Medical Devices via the MDIS.
Documentation requirements: Application and local-responsibility records; accepted QMS evidence; technical documentation in the required STED format; classification rationale; labeling/IFU samples; and the applicable conformity evidence. Use the accepted reference-market approval documents or an MDACS CAB certificate as appropriate, together with the required Essential Principles declaration/checklist.
Timeline: vetting and approval of a listing application should be completed within 12 weeks of submitting the application, after all required supporting information is available; applicant questions and dossier readiness can extend total calendar time.
Renewal: Listing approval (including conditional approval) is valid for five (5) years. Renewal applications should be submitted between 12 weeks and 1 year before expiry; if the listing expires, a new application is required.
Expedited Approval Scheme (selected Class II/III/IV general medical devices; Class B/C/D IVDs)
MDD offers an Expedited Approval Scheme intended to streamline review for eligible MDACS listing applications.
Criteria: No reported deaths/serious injury; no active recalls or field safety corrective actions; and two or more valid reference market approvals.
Process: the LRP applies for inclusion into the List of Medical Devices via the MDIS.
Documentation requirements: The applicable application, QMS, technical, classification and labeling records, together with evidence of the required reference-market approvals and the scheme-specific safety declaration. Verify the current expedited criteria for the actual product and approvals.
Timeline: There is no official turnaround time but applications are reviewed on a priority basis.
Renewal: Listing approval is valid for five (5) years. Renewal applications should be submitted between 12 weeks and 1 year before expiry; if the listing expires, a new application is required.
Other MDACS listing requirements (Hong Kong)
Local Responsible Person (LRP): Mandatory for MDACS device listing submissions for manufacturers with no presence/entities in Hong Kong. The LRP is the applicant of record and must support MDD requests for additional information and inspection of originals/certified copies.
No government application fee: MDD states no fee is charged for inclusion of devices into the MDACS lists (companies should still plan for third-party or operational costs).
Labelling and IFU package: Labeling and IFU can be in English, Chinese, or both. Devices intended for home or lay-use must be labeled in English and Chinese.
Post-market obligations: LRPs must report adverse events within defined timeframes. Device tracking to the user level is required for certain device types.
What documentation is required to list a medical device or IVD in Hong Kong?
Documentation requirements vary by device type (general vs IVD), risk class, and whether you pursue the standard route or the expedited scheme. However, a complete package commonly includes:
Technical documentation in STED format
Evidence demonstrating conformity to Essential Principles, which includes a CAB review certificate or marketing authorization documents from an approved reference market
Labeling samples and IFU
A clear classification rationale per MDACS rules
QMS evidence aligned to ISO 13485
For the Expedited Approval Scheme, documentation supporting the eligibility criteria and the required set of independent regulatory approvals
What is a Hong Kong Local Responsible Person (LRP) and why do you need one?
The Local Responsible Person (LRP) is the Hong Kong-based entity that serves as the official applicant for MDACS device listing. The LRP serves as the primary regulatory liaison between the manufacturer and the MDD. An LRP is required for foreign manufacturers seeking registration; local manufacturers can also appoint an LRP to facilitate the registration process.
The LRP submits and maintains device listing applications, ensures effective communication with users, distributors, and authorities, and maintains comprehensive supply and traceability records for at least the device lifetime or seven years, whichever is longer. They must operate a documented complaint handling system, arrange maintenance and service support where applicable, and implement tracking systems for specified high-risk devices, including annual surveillance reporting. The LRP is also responsible for reporting adverse events within defined timelines, notifying the MDD of recalls and field safety notices, managing corrective actions, and making records available for inspection upon request.
Current listing and procurement decision record
| Question | Evidence to keep |
|---|---|
| MDACS eligibility | Covered device/IVD class and current guidance version |
| Conformity route | Accepted reference evidence or applicable CAB assessment for the actual device |
| Expedited eligibility | Current scheme conditions and independent approvals, not merely two certificates of any kind |
| Local responsibilities | LRP appointment, document access, safety reporting and renewal owner |
| Procurement | The actual tender's requirements and Department of Health Stage C rules where applicable |
The MDACS overview and activities register record current guidance and updates. Department of Health Stage C procurement measures took effect March 23, 2026; assess that scope separately from the voluntary listing framework. Listing is not an unrestricted approval of every claim or a replacement for other applicable controls.
 Regulatory Update · Oct 1, 2026ANVISA–Hong Kong DH Cooperation MoURegulatory Update · Sep 5, 2026Hong Kong MDACS GN Update (Sep 2026)Regulatory Update · Aug 20, 2026Hong Kong GN-02 and GN-06 Listing NotesRegulatory Update · Aug 20, 2026Hong Kong GN-10 Non-reportable ChangesBlog · Jul 13, 2026Aesthetic Devices: The 2026 Global Market-Access MapBlog · May 29, 2026Turn FDA & EU Approvals Into Global Market AccessVideo · Oct 16, 2026Hong Kong MDACS: Voluntary ListingVideo · Aug 11, 2025How to register a medical device in Hong KongCase Study · Aug 5, 2026Vietnam IVD Registration for an Oncology AssayBlog · Oct 5, 2026China GMP inspection preparation
Faster submissions. Predictable costs. Local support.
We provide turnkey regulatory support in Hong Kong, all for a flat annual fee:

Dossier preparation and submission, powered by AI assisted workflows and regulatory experts.
In country representation, with Pure Global serving as your Local Responsible Person.
Post market surveillance support, including vigilance reporting and authority communications.
Importer authorizations, post approval modifications, and renewals.
Frequently asked questions
Pure Global offers flat-fee annual pricing for bundled medical device and IVD registration and representation in Hong Kong, starting at $2,000/year. The flat fee includes dossier preparation and submission, in-country representation, translation, modifications, distributor authorization, and post-market support. The fee does not include certified translations or translations from languages other than English.
Flat fee structure for Class I/II devices (Class A/B IVDs):
1 device = $2,000/year
2-5 devices = additional $1,000 per device per year (e.g., 4 devices = $5,000/year)
6-10 devices = additional $500 per device per year (e.g., 8 devices = $7,500/year)
Flat fee structure for Class III/IV devices (Class C/D IVDs):
1 device = $3,000/year
2-5 devices = additional $1,500 per device per year (e.g., 4 devices = $7,500/year)
6-10 devices = additional $1000 per device per year (e.g., 8 devices = $12,000/year)
Contact us for a custom quote if you have 11+ devices.
A three-year contract is required to lock in the fees quoted above. However, other contractual arrangements are available. Use our Fee Calculator for an instant estimate and more information about flat fee terms.
Medical devices in Hong Kong are classified from Class I-IV; IVDs are classified from A-D. Understanding the classification of your device is essential for determining the appropriate regulatory pathway. The voluntary registration system specifically targets Class II or higher medical devices and Class D IVD products.
No, the registration system for medical devices in Hong Kong is voluntary, particularly under the Medical Device Administrative Control System (MDACS). However, the Department of Health encourages the registration of Class II or higher medical devices and Class D IVD products as soon as possible under Hong Kong medical device regulations.
Manufacturers without a registered business in Hong Kong must appoint a Local Responsible Person (LRP). The LRP is responsible for registering the products with the Medical Device Division (MDD Hong Kong) and fulfilling post-market surveillance responsibilities.
Single Process,
Multiple Markets
When you partner with Pure Global, a single registration process opens doors to multiple countries. Our global subsidiaries make this streamlined path possible.








Let's Talk,
Anywhere You Are.
Whether looking for more information or ready to partner with us, we're here to guide you through every step of the regulatory process.
Contact us