TFDA Taiwan Medical Device Registration
Last reviewed:
Plan Taiwan medical device access through the applicable TFDA pathway, with local responsibilities, product evidence, labeling and lifecycle controls defined before launch.
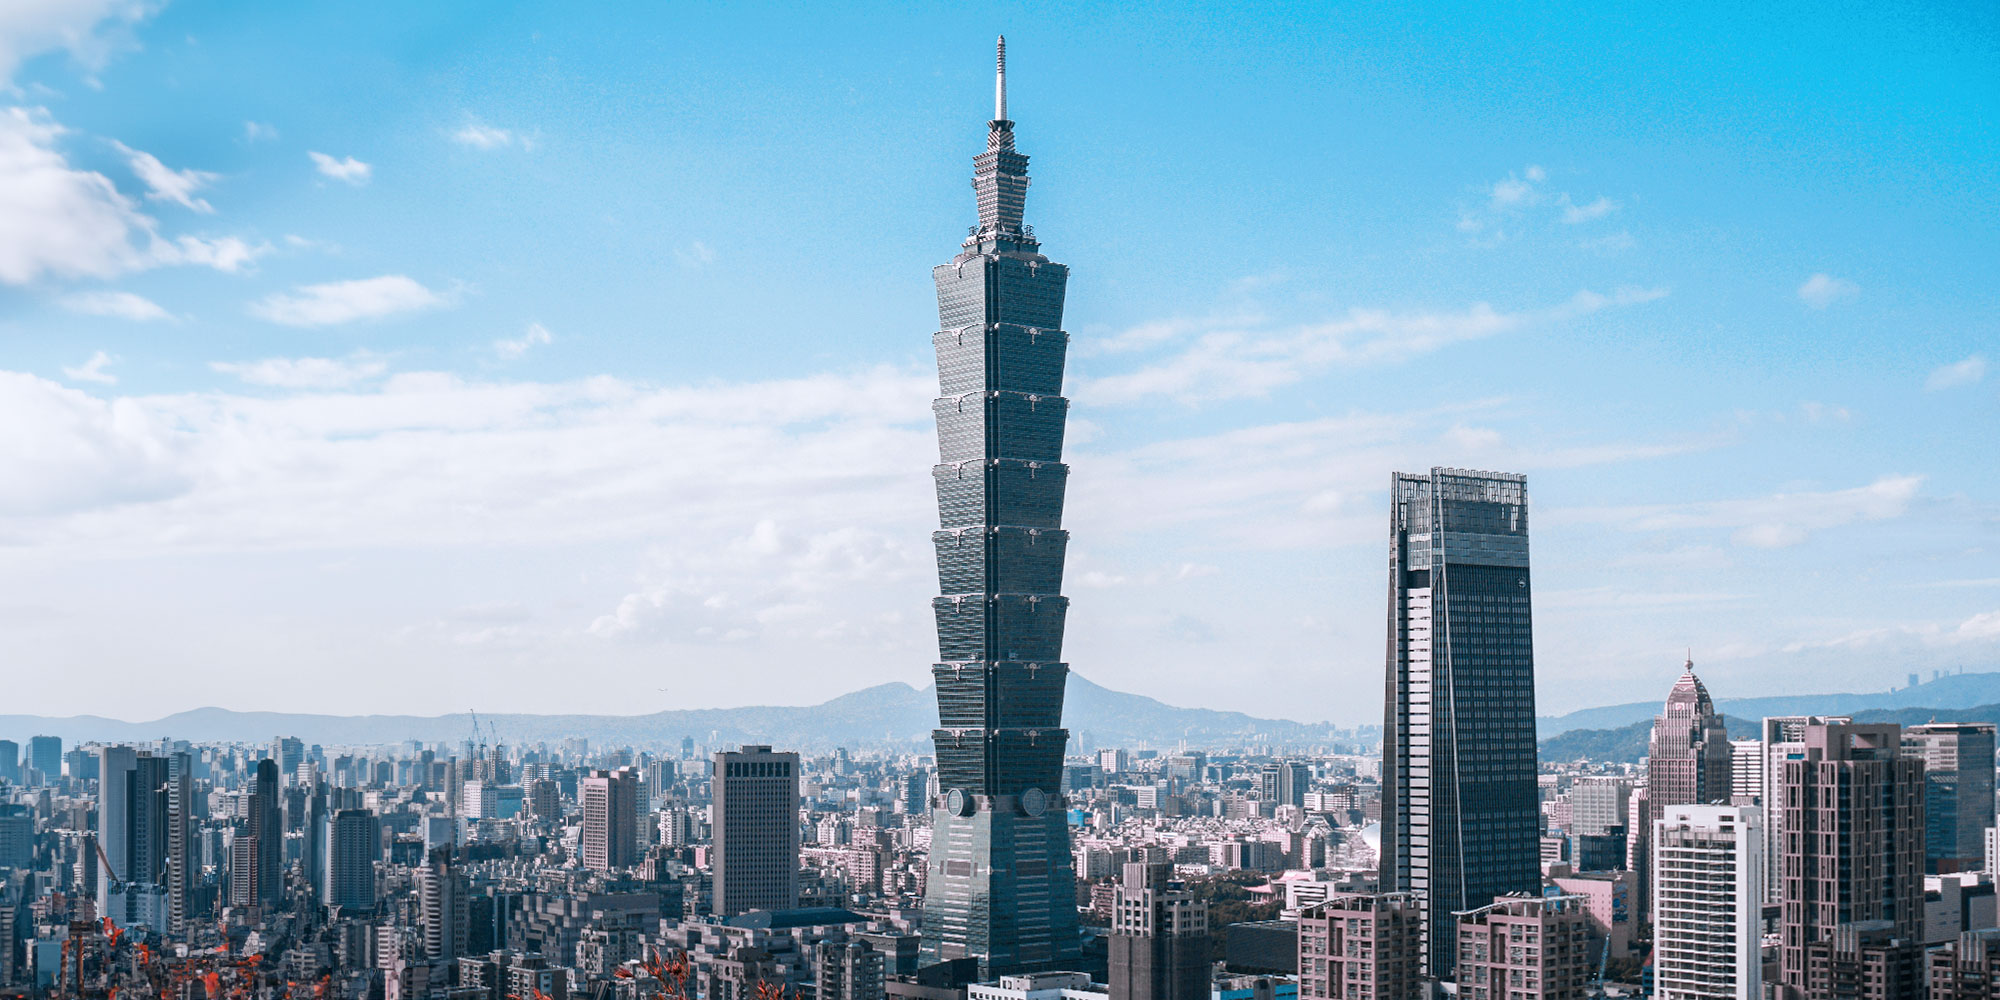
Navigating the TFDA's Requirements
Medical devices and in vitro diagnostics in Taiwan are regulated by the Taiwan Food and Drug Administration (TFDA) under the Ministry of Health and Welfare. The Medical Devices Act provides the central legal framework. TFDA maintains current rules and guidance on its official medical devices page.
Taiwan medical device classification
Taiwan uses Classes 1, 2, and 3, from lower to higher risk. Classification is based on the intended use and product characteristics and is assigned against Taiwan's classification regulations and product categories. A US, EU, or other overseas class can inform the analysis but does not determine the Taiwan result.
Lower-risk Class 1 products may follow a listing route, while Class 2 and Class 3 products generally require registration and a fuller technical review. The exact pathway, evidence, and whether a simplified route applies depend on the product category, prior approvals, and current TFDA rules. IVDs should be assessed under the classifications and requirements applicable to their intended testing purpose.
Local license holder and QMS review
A foreign manufacturer generally needs a Taiwan-based medical device business to act as the local license holder and regulatory contact. The agreement should cover dossier access, authority responses, change reporting, license maintenance, complaints, vigilance, recalls, and transfer arrangements.
Quality-system compliance is a separate workstream from the product application. Depending on the device and manufacturing activities, the manufacturer may need TFDA quality-management-system documentation review, often referred to in practice as QMS or historically QSD review. Exemptions or simplified evidence for particular lower-risk products should be confirmed rather than assumed.
Technical file and lifecycle
A registration file commonly includes intended use, classification, device descriptions, manufacturer and site information, authorization documents, labeling and instructions for use, quality-system evidence, risk management, verification and validation, and clinical or performance evidence proportionate to risk. Chinese labeling and the local license-holder details must match the approved product.
After listing or registration, the manufacturer and license holder must keep product, site, quality certificates, and label information current. Complaints, adverse events, field safety corrective actions, and recalls need coordinated procedures. Design, software, intended-use, site, or labeling changes should be assessed before implementation to determine whether TFDA notification, approval, or a new application is required.
Taiwan pathway and evidence decisions
| Workstream | Local requirement or decision |
|---|---|
| Product class | Local classification under the Medical Devices Act framework |
| Class 1 | Applicable listing route and product-specific exceptions |
| Class 2/3 | Registration with appropriate technical/evidence review |
| Manufacturing system | Separate QMS/QSD assessment, exemptions and accepted evidence |
A product licence and manufacturing quality-system review are different records. Do not present a commercial ISO 13485 certificate as the same thing as TFDA QMS acceptance, or claim one exemption for every nonsterile Class 1 device without checking the applicable activities and rules.
Use the official TFDA instructions and related official process guidance for the filing scope. Maintain a controlled record of the chosen route, current evidence, open gaps, responsible parties and authority responses. Separate fees, document preparation, review and applicant work when planning the launch.
For a multi-country program, use the market directory to coordinate evidence reuse while retaining the local applicant, language, registration and postmarket requirements for each destination.
 Regulatory Update · Oct 17, 2026TFDA Clinical Trial E-SubmissionsRegulatory Update · Sep 14, 2026Taiwan Recall Rework AmendmentRegulatory Update · Sep 14, 2026Taiwan TFDA Filing and Label ApprovalBlog · Jul 27, 2026IVD Global Market Access 2026Blog · May 29, 2026Turn FDA & EU Approvals Into Global Market AccessRegulatory Update · Oct 27, 2026Pakistan DRAP PSW Device Import RulesCase Study · Aug 5, 2026Vietnam IVD Registration for an Oncology AssayBlog · Oct 5, 2026China GMP inspection preparationBlog · Sep 6, 2026One Device, Three UDI DatabasesVideo · Oct 16, 2026South Korea MFDS: Device Registration
Tailored Support for Market Entry
Our local partner’s expertise in Taiwan's medical device market offers you:

Assistance with product classification and identification of the regulatory pathway.
Compilation of the technical dossier required for TFDA registration.
Acting as your Taiwan Agent, managing the registration with TFDA and ensuring compliance with post market regulations.
Frequently asked questions
No. Assess TFDA’s manufacturing quality-system requirements, accepted evidence and applicable exemptions separately. A commercial quality certificate and the local QMS/QSD outcome are not interchangeable.
No. Confirm the specific category, listing eligibility, exemptions and local-business requirements. Product classification and manufacturing-system applicability require separate documented assessments.
Define regulatory access, product/site scope, authority responses, changes, licence and quality-record maintenance, safety reporting, recalls and transfer arrangements. Keep the approved Chinese product information aligned with the evidence.
Single Process,
Multiple Markets
When you partner with Pure Global, a single registration process opens doors to multiple countries. Our global subsidiaries make this streamlined path possible.








Let's Talk,
Anywhere You Are.
Whether looking for more information or ready to partner with us, we're here to guide you through every step of the regulatory process.
Contact us