COFEPRIS Mexico Medical Device Registration
Last reviewed:
Plan COFEPRIS medical device registration with the correct class, standard or eligible abbreviated route, Mexico registration holder, Spanish labeling and lifecycle controls.
How to register a medical device in Mexico
Medical device and In Vitro Diagnostic (IVD) device registrations in Mexico are regulated by COFEPRIS (Comisión Federal para la Protección contra Riesgos Sanitarios). Registration requirements are established under the General Health Law (Ley General de Salud) and the Regulations of Health Supplies (Reglamento de Insumos para la Salud), alongside applicable NOM standards such as NOM-241-SSA1-2025 for Good Manufacturing Practices.
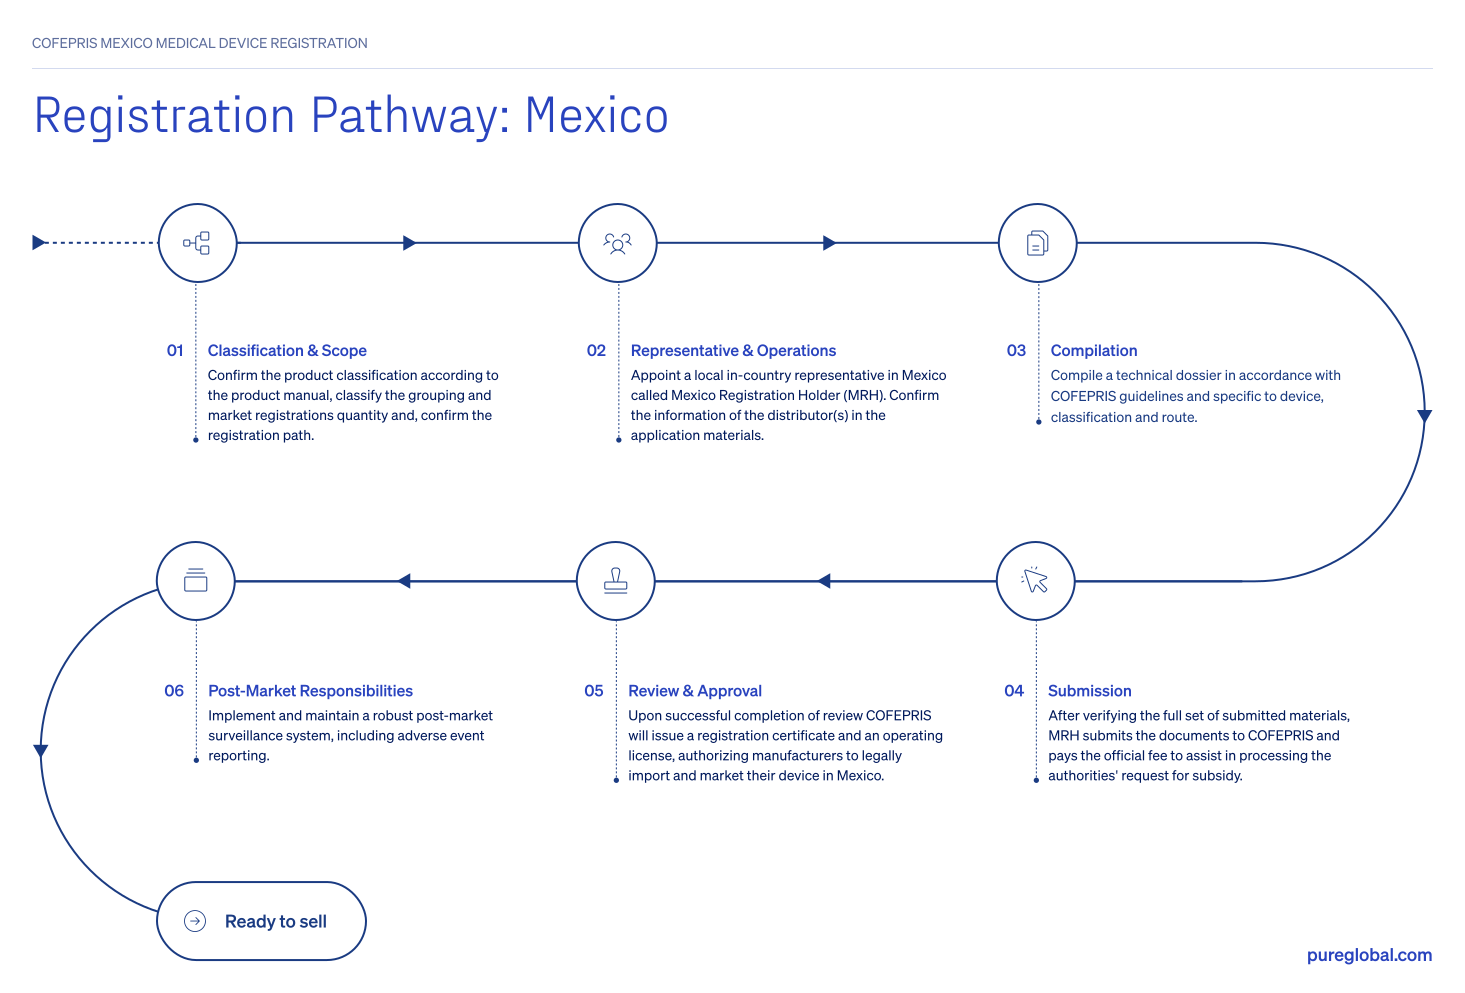
Foreign manufacturers must appoint a local Mexico Registration Holder (MRH). The MRH is a legally established entity in Mexico who acts as the official representative for the foreign manufacturer. The MRH is responsible for obtaining and holding the sanitary registrations for your medical devices, as well as coordinating with COFEPRIS. It is possible to transfer your registrations to a different holder through formal regulatory channels. However, this process can be cumbersome, so it’s important to thoroughly vet your chosen MRH. Once appointed, the MRH will identify which regulatory pathways, depending on the product and available reference approvals, the foreign Manufacturer shall follow.
COFEPRIS Mexico medical device regulatory pathways
Mexico offers two registration routes: the Standard Route and the Equivalency (Abbreviated) Route:
Standard Route (Full Technical Review)
Process: COFEPRIS conducts a comprehensive review of the submitted technical dossier, including clinical, safety, and labeling documentation.
Requirements: A complete technical file with clinical and safety data, quality system evidence, labeling in Spanish, a Free Sale Certificate or Foreign Government Certificate, and a Letter of Representation.
Timeline: Review timelines vary due to the complexity and volume of submissions.
Renewal: Initial registrations are valid for 5 years; under the January 2026 health-law reform — extended by the April 2026 regulations reform to first and subsequent extensions alike — renewals are granted in 10-year terms.
Equivalency Route (Abbreviated Pathway)
Process: COFEPRIS grants expedited authorization based on prior approval from a recognized reference authority.
Eligibility: Under the Lineamientos published in the DOF on July 18, 2025 (applications opened September 1, 2025), device eligibility rests on prior authorization from a regulator of a country on the IMDRF Management Committee — covering FDA, EU CE marking, Health Canada, Swissmedic, ANVISA, TGA, MFDS, and NMPA — with full MDSAP members additionally recognized for GMP certification. The ICH- and WHO-listed authorities in the Lineamientos apply to medicines, not devices.
Requirements: The device must be identical to the version approved by the reference authority. Required documentation includes a Certificate of Free Sale and conformity assessment, foreign approval, Clinical Evaluation Report, risk analysis, draft labeling in Spanish, and quality system evidence.
Timeline: COFEPRIS should issue a decision within 30 business days of receiving a complete application. Delays may occur when there is a high volume of submissions.
Renewal: Initial registrations are valid for 5 years; renewals may be granted for up to ten years under the 2026 Article 376 framework, subject to the applicable requirements and issued term.
Other COFEPRIS registration requirements
Mexico Registration Holder (MRH): Mandatory for all foreign manufacturers. The MRH submits applications, pays fees, and acts as the local regulatory contact with COFEPRIS.
Distributor information: Distributor details must be confirmed and included in the application materials.
Translation: All submissions and labeling must be in Spanish.
Quality system evidence: While ISO 13485 is not legally required, COFEPRIS expects evidence of a robust quality management system, under the applicable route. Assess accepted GMP certification or equivalent evidence; a CE mark alone should not be treated as a universal GMP certificate.
Post-market obligations: Manufacturers must maintain post-market surveillance systems, including adverse event reporting, throughout the registration lifecycle.
What documentation is required to register a medical device or IVD with COFEPRIS?
Documentation requirements depend on the device classification and registration route, but a complete dossier generally includes:
Free Sale Certificate or Foreign Government Certificate (from reference market if using Equivalency route)
Certificate of Good Manufacturing Practices, ISO 13485, CE Mark, MDSAP, etc.
Foreign approval (required for Equivalency route)
Technical file, including clinical and safety data (required for Standard route)
Device labeling in Spanish
Letter of Representation
What is a Mexico Registration Holder and why do you need one?
Before submitting a medical device or IVD registration to COFEPRIS, foreign manufacturers must appoint a Mexico Registration Holder (MRH) to serve as their local regulatory representative. The MRH is responsible for submitting registration dossiers, paying government fees, and maintaining communication with COFEPRIS throughout the product’s approval and post-market lifecycle.
Sanitary registrations in Mexico are issued in the name of the MRH. This means the MRH is legally recognized as the registration holder. Because transferring a registration to a new MRH typically requires a formal process with COFEPRIS, manufacturers should evaluate potential representatives carefully before proceeding.
Do foreign medical device manufacturers also need an importer and a distributor in Mexico?
Yes. Foreign manufacturers typically need an MRH, an authorized importer, and distributor to distribute medical devices and IVDs in Mexico. The MRH acts as the legal regulatory representative and holds the sanitary registration issued by COFEPRIS, while the importer and distributor are responsible for physically bringing the product into Mexico and ensuring compliance with customs and labeling requirements. These roles can be fulfilled by the same organization or by separate entities, but all must be documented and authorized as part of the regulatory process.
A Mexico launch and maintenance evidence map
| Workstream | Decision to document | Next guide |
|---|---|---|
| Product class and grouping | Mexican rules, low-risk list and actual model family | Classification |
| Abbreviated route | Eligible reference authorization, identical device and required evidence | Regulations |
| Labeling transition | Current NOM-137-SSA1-2008 and preparation for the final 2025 successor | Labels and IFU |
| Imports and lifecycle | Holder/importer responsibilities, permits, changes and reporting | Postmarket compliance |
The 2026 health-law reform and final NOM-137-SSA1-2025 have different functions and effective-date provisions. The new labeling standard's 360-day period must not be described as if every new requirement is already mandatory. Keep product authorization, GMP evidence and importer permissions separate, and confirm the actual renewal decision rather than assuming an automatic ten-year extension.
 Case Study · Aug 5, 2026COFEPRIS IVD Registration for a Reagent MenuRegulatory Update · Jul 8, 2026Mexico COFEPRIS Registration Guidance 2026Regulatory Update · May 26, 2026Mexico NOM-137-SSA1-2025 Medical Device Labeling UpdateRegulatory Update · Mar 25, 2026COFEPRIS 2026 Guidance on Electronic NotificationsRegulatory Update · Mar 25, 2026COFEPRIS 2026 Modifications Guideline for Medical DevicesBlog · Aug 24, 2026Medical Device Registration Verification 2026Blog · Jun 17, 2026How to register medical devices and IVDs in MexicoBlog · May 29, 2026Turn FDA & EU Approvals Into Global Market AccessBlog · Sep 23, 2025Medical Device Registration Costs and Timelines for Brazil, Mexico, and ColombiaVideo · Sep 11, 2026ISO 15223-1: What Device Labels Need to Say
Faster submissions. Predictable costs. Local support.
We provide turnkey regulatory support in Mexico, all for a flat annual fee:

Dossier preparation and submission, powered by AI assisted workflows and regulatory experts.
In country representation, with Pure Global serving as your Mexico Registration Holder.
Post market surveillance support, including vigilance reporting and authority communications.
Adding or changing distributor(s) to the registration, post approval modifications, and renewals.
Frequently asked questions
No. An eligible reference authorization may support the abbreviated route, but the product must meet the route’s identity and evidence conditions and complete the Mexican process with an eligible local registration holder.
The 2026 Article 376 framework permits renewals for up to ten years subject to the applicable requirements. Confirm the issued term and current instructions; the initial registration remains a separate five-year authorization.
It is final and published, but its text provides a 360-day period before entry into force. Maintain current compliance while preparing the successor requirements and assess any inventory transition under the stated conditions.
Use the current official fee category and applicable procedure, then separately budget testing, certifications, translations, local-holder services and import work. Avoid undated currency conversions or a single guaranteed approval timeline.
Single Process,
Multiple Markets
When you partner with Pure Global, a single registration process opens doors to multiple countries. Our global subsidiaries make this streamlined path possible.








Let's Talk,
Anywhere You Are.
Whether looking for more information or ready to partner with us, we're here to guide you through every step of the regulatory process.
Contact us