MHRA UK Medical Device Registration & Approval
Last reviewed:
Plan Great Britain market access through UKCA or an eligible CE-marked route, with MHRA registration, a UK Responsible Person where required and current postmarket controls. Assess Northern Ireland separately.
How to register a medical device in the United Kingdom
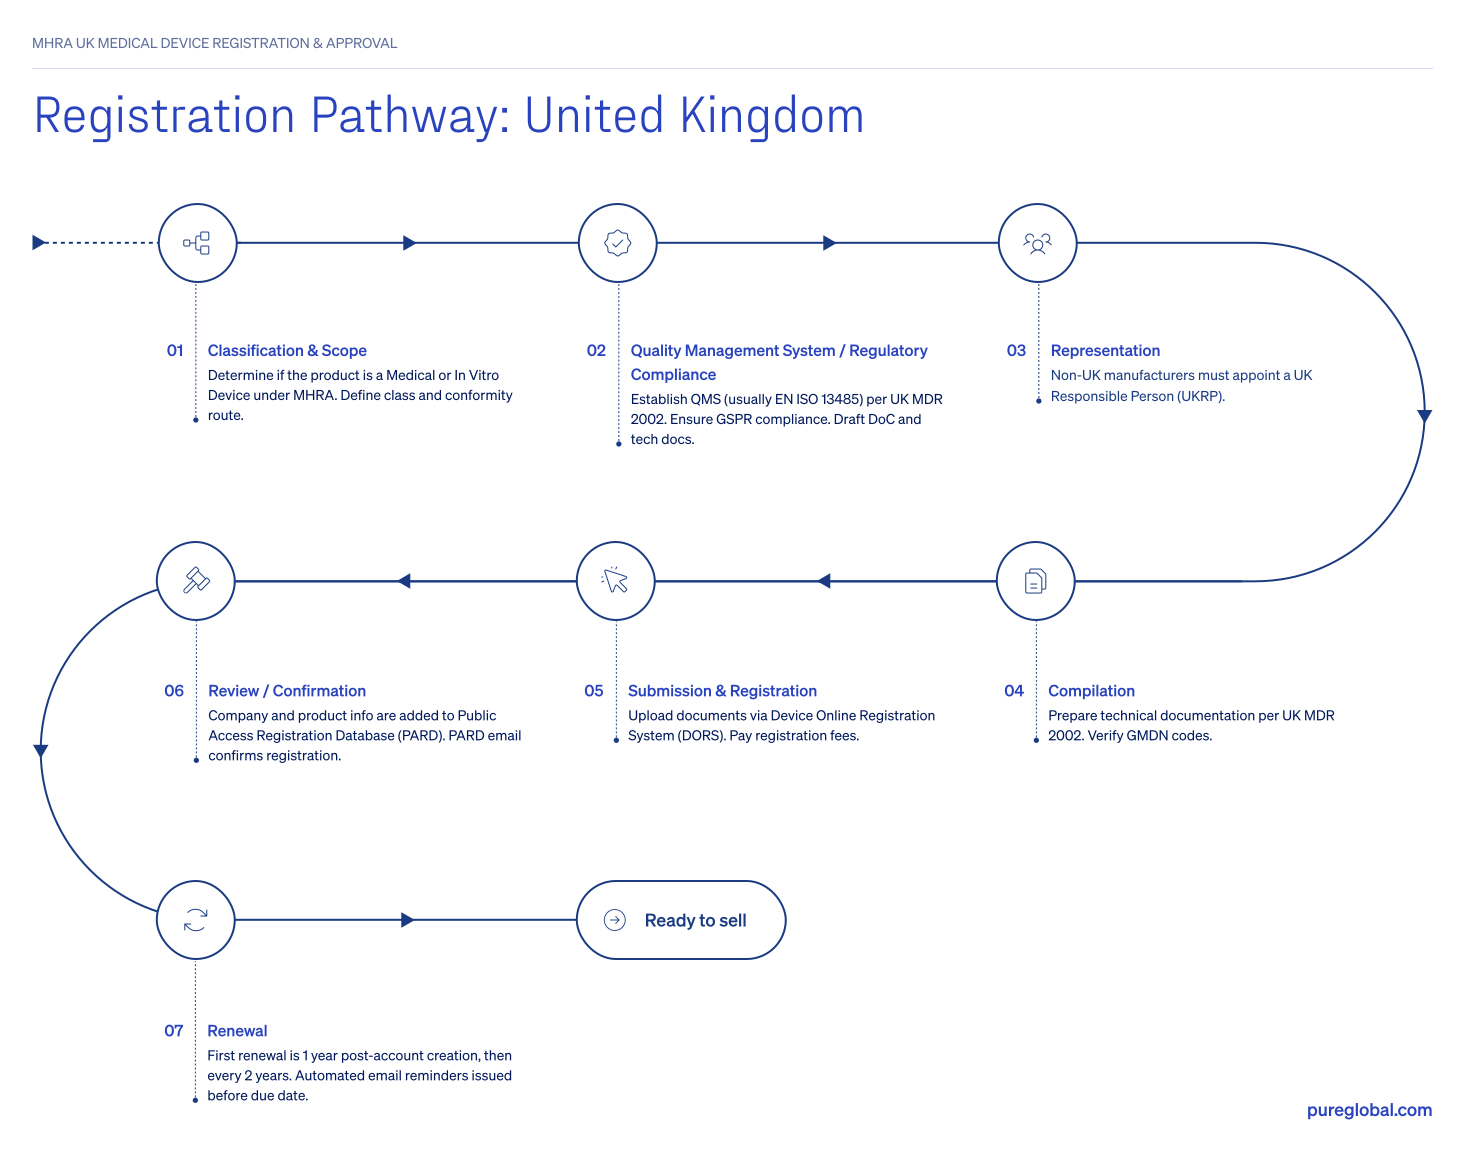
Medical devices and IVDs placed on the market in Great Britain are regulated by the Medicines and Healthcare products Regulatory Agency (MHRA). The primary legislation is the UK Medical Devices Regulations 2002 (UK MDR 2002), and its amendments following the UK’s withdrawal from the European Union.
For Great Britain, devices need the applicable UKCA route or an eligible recognized CE-marked route. CE recognition deadlines vary by the underlying legislation and can extend to 2030; certificate validity and the specific conditions matter. Northern Ireland follows a separate framework. Manufacturers must also register their devices with the MHRA’s Device Online Registration System (DORS).
Foreign manufacturers must appoint a UK Responsible Person (UKRP) if they do not have a UK-based legal presence. The UKRP acts on behalf of the manufacturer to carry out specific regulatory obligations, including device registration in DORS and communications with the MHRA.
United Kingdom medical device and IVD classification
The UK applies a risk-based classification system aligned with EU rules. General medical devices are classified as Class I, IIa, IIb, and III under the UK MDR 2002. Active implantable medical devices follow specific provisions. In vitro diagnostic devices are classified as general IVDs, IVDs for self-testing, and Annex II List A and List B IVDs. Classification determines the conformity assessment route and level of Notified Body involvement.
MHRA medical device regulatory pathways
There are two principal conformity assessment pathways for Great Britain: UKCA conformity assessment, involving a UK Approved Body where required, or eligible CE recognition during the applicable transitional periods. The appropriate route depends on device classification, manufacturer location, and current regulatory timelines.
UKCA marking is not recognized in Northern Ireland, where different authorization rules apply.
UKCA Route (Great Britain market)
Process: The manufacturer completes the applicable UK MDR 2002 conformity assessment. Non-sterile, non-measuring Class I devices and general IVDs can generally use self-declaration; a UK Approved Body is required when the relevant category calls for third-party assessment. Upon successful assessment, the manufacturer affixes the UKCA mark, appoints a UKRP (as applicable) and registers the device in DORS.
Requirements: Documentation includes a technical file or design dossier demonstrating conformity with the Essential Requirements, a Declaration of Conformity, evidence of an appropriate quality management system (typically certified to ISO 13485), clinical evidence where applicable, and labeling compliant with UK requirements.
Timeline: Review timelines depend on device classification and the scope of UK Approved Body assessment. Higher-risk devices will have longer assessment periods.
Renewal: UKCA certificates issued by a UK Approved Body are typically valid for up to five years. DORS must be maintained and updated as required.
CE Marking Route (Recognized during transitional periods)
Process: The manufacturer completes conformity assessment under applicable EU legislation with an EU Notified Body where required. During recognition periods established by UK law, CE-marked devices may be placed on the UK market. To do so, the manufacturer shall appoint a UKRP (as applicable) and must register the CE-marked devices in DORS.
Requirements: Required documentation includes a technical dossier that meets EU requirements, an EU Declaration of Conformity, applicable CE certificates, and compliance with UK registration and labeling provisions, including identification of the UK Responsible Person where applicable.
Timeline: Driven primarily by EU conformity assessment procedures and availability.
Renewal: CE certificates must remain valid under EU rules. Continued placement on the Great Britain market depends on compliance with applicable UK transitional provisions.
Key MHRA registration requirements
UK Responsible Person: Required for manufacturers established outside the UK placing devices on the Great Britain market. The UK Responsible Person must be established in the UK and is responsible for registering devices with the MHRA, maintaining technical documentation access, and facilitating communication with MHRA as applicable
Registration with MHRA: All medical devices and IVDs must be registered with the MHRA before being placed on the Great Britain market. Registration is completed through the MHRA Device Online Registration System (DORS). Annual fees apply.
Labeling: Devices must bear the UKCA or CE Mark as applicable. Where a UKRP is required and UKCA marking is affixed, its name and address must appear on the product labeling, outer packaging or IFU as permitted by the applicable rules. UKRP details do not need to be included on labelling for CE marked devices, unless the device bears both the CE and UKCA markings. Instructions for Use and labeling must be in English.
Post-market obligations: Manufacturers, with the support of their UKRP, must establish post-market surveillance systems, report serious incidents and field safety corrective actions to the MHRA, and comply with vigilance requirements set out in the UK MDR 2002.
What documentation is required to register a medical device or IVD in Great Britain?
Documentation requirements depend on device classification and the selected conformity assessment route. In general, manufacturers should prepare:
A complete technical documentation file demonstrating conformity with the Essential Requirements of the UK MDR 2002 for the UKCA marked device. If leveraging CE marking, then a complete technical documentation file demonstrating conformity with the applicable EU legislation is required.
A Declaration of Conformity referencing applicable UK legislation for UKCA marked devices. If leveraging CE marking, the Declaration of Conformity must reference applicable EU legislation.
Quality management system certification, where required.
Clinical evaluation or performance evaluation documentation, as applicable.
UKCA or CE certification from an Approved Body or EU Notified Body, where required.
Pay the annual fees once the MHRA device registration is submitted. MHRA fees will be charged based on the MHRA annual fee period.
What is a UK Responsible Person and why do you need one?
A UK Responsible Person (UKRP) is a legal entity established in the United Kingdom that acts on behalf of a manufacturer located outside the UK. The UK Responsible Person ensures that the Declaration of Conformity and technical documentation have been prepared, registers the device with the MHRA, cooperates with the MHRA on corrective actions, and maintains copies of technical documentation for inspection. Non-UK manufacturers cannot place devices on the Great Britain market without appointing a UKRP.
What is a UK Approved Body?
A UK Approved Body is an organization designated by the UK government to perform conformity assessments under the UK MDR 2002. Approved Bodies review quality management systems, technical documentation, and clinical evidence for higher-risk devices before issuing UKCA certificates. Only Approved Bodies with an appropriate scope of designation may assess specific device types.
What are the deadlines for leveraging EU CE Marking in Great Britain?
CE Marked devices can be placed on the Great Britain market up until these deadlines:
EU MDD or EU AIMDD general medical devices: earlier of certificate expiry or 30 June 2028
EU IVDD IVDs: earlier of certificate expiry or 30 June 2030
EU MDR general medical devices (including custom-made): 30 June 2030
EU IVDR IVDs: 30 June 2030
Self-declared Class I devices under EU MDR: 30 June 2030
Self-declared Class I devices under MDD (declared before 26 May 2021 and upclassified under MDR): 30 June 2028
Class I sterile or measuring devices with valid MDD certificate: 30 June 2028
Not eligible for the above timelines:
Class I self-certified medical devices under the EU directives that do not meet the EU transitional timelines.
General IVDs under the EU directives that do not meet the EU transitional timelines.
Custom-made devices compliant with EU MDD or EU AIMDD
Devices placed on the Great Britain market after these deadlines must comply with the UK conformity assessment process and bear the UKCA mark (unless a new legislative extension is introduced). Devices lawfully placed on the Great Britain market before the deadline may continue to be made available or put into service. However, any new placement after the deadline requires UKCA compliance.
Great Britain and Northern Ireland require separate plans
| Territory/task | Requirement to assess | Evidence to keep |
|---|---|---|
| Great Britain conformity | UKCA or eligible CE recognition under current rules | Declaration, certificate scope and applicable recognition end date |
| Foreign manufacturer in GB | UK Responsible Person and MHRA registration | Written designation, device scope and account records |
| GB postmarket system | Requirements effective June 16, 2025 | PMS plan, applicable reports and vigilance procedures |
| Northern Ireland | Applicable EU framework and current NI arrangements | EU conformity, representation and registration applicability |
Use the current MHRA framework, registration guidance and PMS implementation notice. A policy proposal or consultation must not be presented as a replacement for the enacted CE-recognition deadlines. Coordinate the UKRP with EU representation and Swiss requirements where the launch spans those territories.
 Regulatory Update · Oct 17, 2026MHRA DORS Annual FeesRegulatory Update · Feb 4, 2026UK MHRA Medical Device Registration Update 2026: New FeesRegulatory Update · Dec 17, 2025Singapore HSA–UK Fast Track Pathway 2025Regulatory Update · Oct 22, 2025UK MHRA 2026 Guidance on Medical Device Registration FeesBlog · Sep 6, 2026One Device, Three UDI DatabasesBlog · Aug 17, 2026Coronary Stents: Eight Highest-Tier FilesBlog · Aug 10, 2026Surgical Robots: Eight Markets, Eight Regulatory FilesBlog · Aug 3, 2026ISO 10993-1:2025: One Core, Seven Regulatory SystemsVideo · Oct 16, 2026MHRA: UK Medical Device RulesVideo · Oct 16, 2026ISO 10993-1:2025: What Changed
Tailored Support for Market Entry
Our services are designed to streamline your regulatory process in the UK:

Assistance with product classification and identification of the UK regulatory pathway.
Compilation of technical documentation required for UKCA marking and MHRA registration.
UK Responsible Person (UKRP) service includes MHRA portal registration and more for an annual flat fee.
Frequently asked questions
From April 1, 2026, the MHRA medical device registration fee is £300 per year for each Level 2 GMDN category, or Level 1 where no Level 2 exists. Registrations added during the charging year are assessed pro rata, and the unit fee can change annually. Confirm the live account categories and current MHRA fee guidance before payment.
Government fees do not include consulting, technical documentation, conformity assessment, or UK Responsible Person services.
A manufacturer outside the UK must appoint a UK Responsible Person for Great Britain, meet the applicable UK MDR 2002 requirements, and register devices with the MHRA before placement on the market. The product may use UKCA or an eligible CE-marked transition route, depending on its certification and the current recognition deadline.
Pure Global offers flat-fee annual pricing for bundled UK Authorized Representative services and regulatory support, starting at $2,000/year. The flat fee includes document review, MHRA portal registration, and Certificate of Free Sale (CFS) processing. Government and other third party fees, when applicable, are not included.
Flat fee structure for medical devices and IVDs:
1 device or device group = $2,000/year
2-5 devices/groups = additional $500 per device per year (e.g., 5 devices = $4,000/year)
6-10 devices/groups = no additional cost (e.g., 8 devices - $4,000/year)
Contact us for a custom quote if you have 11+ devices.
A three-year contract is required to lock in the fees quoted above. However, other contractual arrangements are available. Use our Fee Calculator for an instant estimate and more information about flat fee terms.
Single Process,
Multiple Markets
When you partner with Pure Global, a single registration process opens doors to multiple countries. Our global subsidiaries make this streamlined path possible.








Let's Talk,
Anywhere You Are.
Whether looking for more information or ready to partner with us, we're here to guide you through every step of the regulatory process.
Contact us