DRAP Pakistan Medical Device Registration
Last reviewed:
Plan Pakistan medical device access through the applicable DRAP pathway, with local responsibilities, product evidence, labeling and lifecycle controls defined before launch.
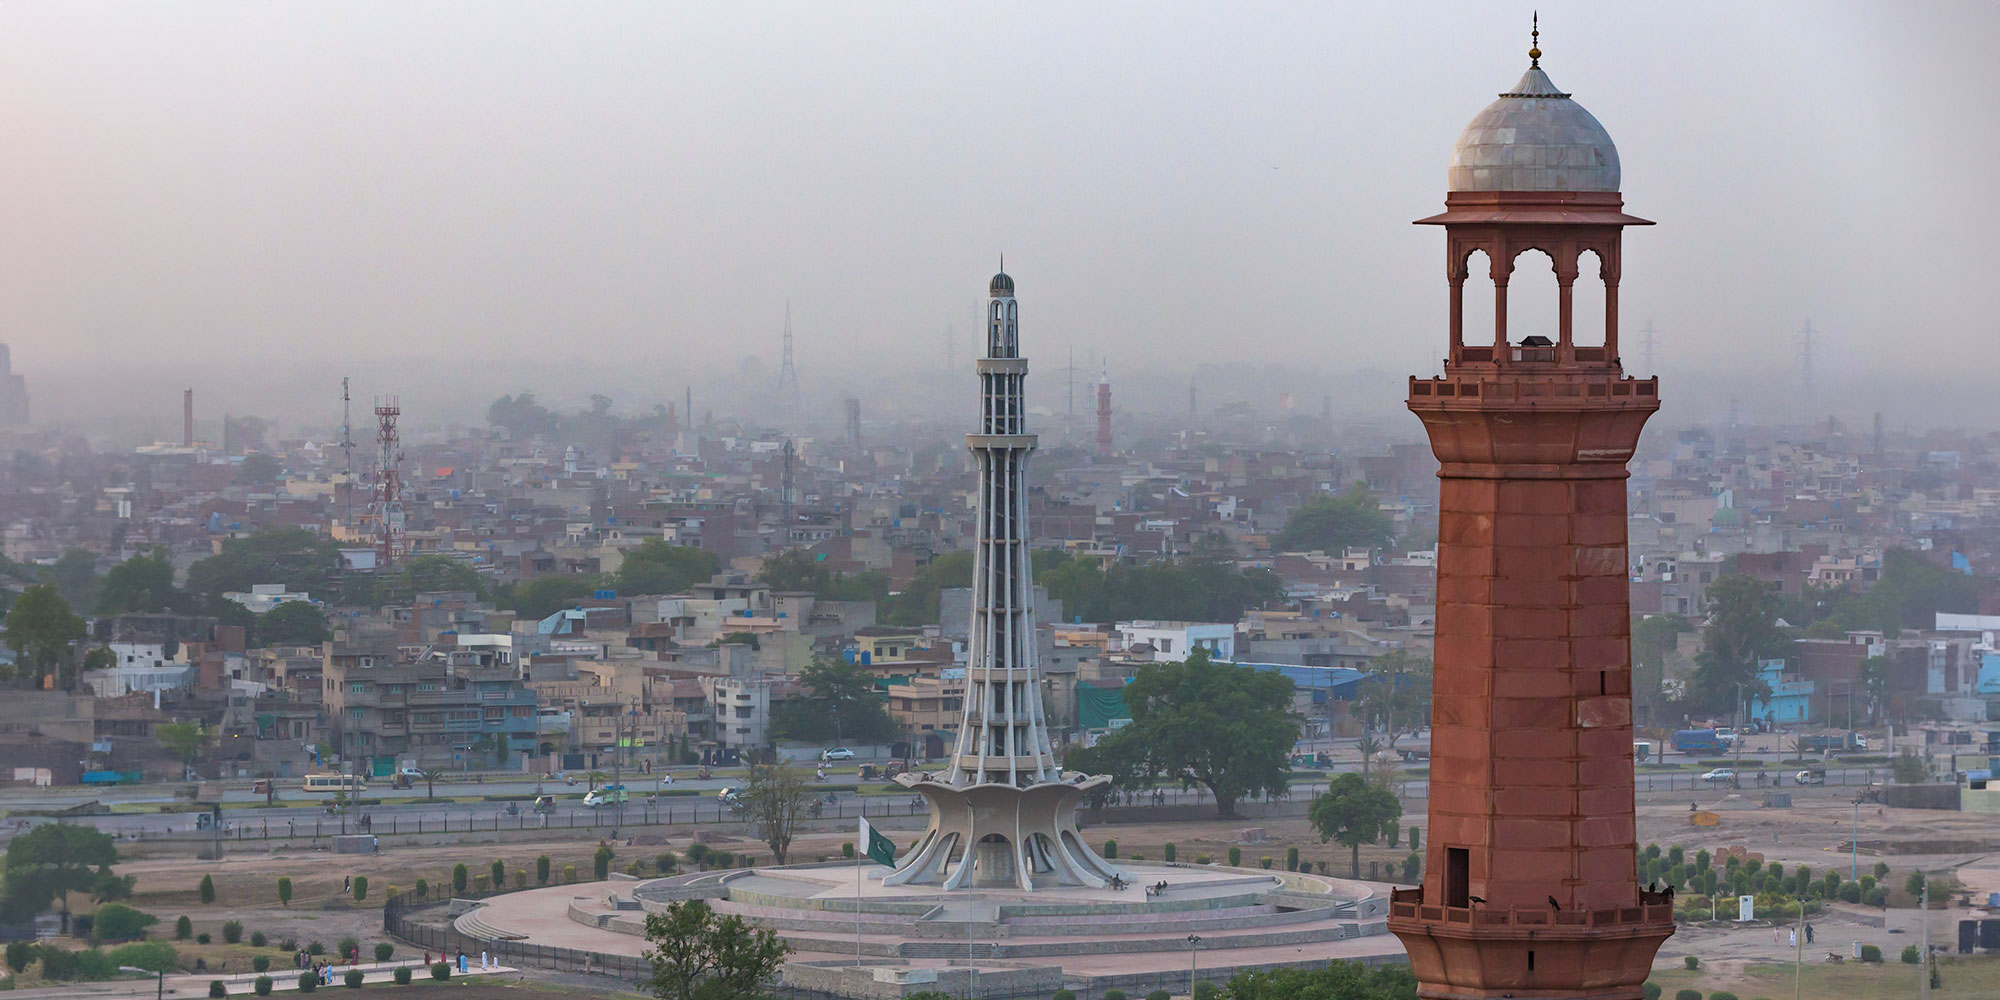
Complying with DRAP Regulations
Medical devices in Pakistan are regulated by the Drug Regulatory Authority of Pakistan (DRAP) under the Medical Devices Rules, 2017 and subsequent amendments. DRAP oversees establishment licensing, product applications, import controls, market surveillance, and safety actions. Manufacturers should use the official DRAP medical device application guidance to confirm the current forms, fees, and portal requirements before filing.
Pakistan medical device classification
Pakistan uses four risk classes: Class A for the lowest-risk devices through Class D for the highest-risk devices. Classification depends on intended use, duration of contact, invasiveness, whether the device is active, and the consequences of a failure. The class determines the application route and the depth of technical, clinical, and quality-system evidence DRAP expects. IVDs also follow a risk-based framework, so the correct classification should be documented before a submission is assembled.
Local establishment and applicant requirements
A manufacturer outside Pakistan normally works through a locally licensed importer or authorized representative. The Pakistan entity needs the appropriate DRAP establishment license for its activities before it can sponsor product applications or import devices. Manufacturer and site information, authorization letters, quality-system evidence, and proof of approval or free sale from relevant reference markets may be required depending on the product and route.
Commercial control should be addressed in the representation agreement. It should define access to the registration dossier, change-notification responsibilities, complaint and vigilance handling, renewal work, and how registrations can be transferred if the relationship ends.
Registration dossier and lifecycle
A typical file includes device descriptions, intended use, classification rationale, labeling and instructions for use, manufacturing-site details, ISO 13485 evidence, declarations or certificates of conformity, test reports, and clinical or performance evidence where applicable. DRAP may ask questions or request additional documentation during review.
Approval is not the end of the program. Import documentation, approved labeling, distribution records, complaints, adverse-event reporting, field safety corrective actions, and registration or license maintenance must stay aligned. Changes to the manufacturer, design, intended use, labeling, or quality system should be assessed before implementation to determine whether DRAP notification or approval is required.
Pakistan pathway and evidence decisions
| Workstream | Local requirement or decision |
|---|---|
| Importer establishment | Obtain the applicable DRAP establishment licence before the product application |
| Imported Class A | Enlistment route; official forms identify Form 6-A |
| Imported Classes B/C/D | Registration route; official forms identify Form 7-A |
| Domestic manufacture | Separate establishment and product forms; do not copy the import route |
Use the current DRAP digital process and forms rather than a historical paper checklist alone. Confirm the device scope, site, authorization, quality evidence and reference certificates where required. Establishment licensing, product enlistment/registration and shipment readiness are separate milestones.
Use the official DRAP instructions and related official process guidance for the filing scope. Maintain a controlled record of the chosen route, current evidence, open gaps, responsible parties and authority responses. Separate fees, document preparation, review and applicant work when planning the launch.
For a multi-country program, use the market directory to coordinate evidence reuse while retaining the local applicant, language, registration and postmarket requirements for each destination.
 Regulatory Update · Oct 20, 2026Singapore HSA Integration and HTA ScopeRegulatory Update · Oct 17, 2026TFDA Clinical Trial E-SubmissionsRegulatory Update · Oct 17, 2026TGA Digital ScribesRegulatory Update · Oct 11, 2026HSA GN-15 R15 Registration UpdatesCase Study · Aug 5, 2026Vietnam IVD Registration for an Oncology AssayBlog · Oct 5, 2026China GMP inspection preparationBlog · Sep 6, 2026One Device, Three UDI DatabasesBlog · Aug 24, 2026Medical Device Registration Verification 2026Blog · Aug 17, 2026Coronary Stents: Eight Highest-Tier FilesVideo · Oct 16, 2026South Korea MFDS: Device Registration
Tailored Support for Market Entry
Our local partner’s expertise in Pakistan's medical device market offers you:

Assistance with product classification and determination of the regulatory pathway.
Preparation and compilation of the technical dossier required for DRAP registration.
Acting as your Authorized Representative, managing the submission to DRAP and overseeing post market compliance.
Frequently asked questions
No. The official form framework distinguishes Class A enlistment from Class B/C/D registration, including separate imported-product forms. Apply the current process and product scope before preparing the file.
No. DRAP’s application guidance identifies importer establishment licensing as a prerequisite. Confirm the relevant company permissions and product authorization separately before relying on either for supply.
Keep device models, manufacturer/sites, authorized local activities, labeling, quality evidence and import records current. Assess changes and maintain complaint, reporting, recall and renewal responsibilities under the applicable route.
Single Process,
Multiple Markets
When you partner with Pure Global, a single registration process opens doors to multiple countries. Our global subsidiaries make this streamlined path possible.








Let's Talk,
Anywhere You Are.
Whether looking for more information or ready to partner with us, we're here to guide you through every step of the regulatory process.
Contact us