EU IVDR: Classification, CE Marking & Consulting
Last reviewed:
In 2017, the EU introduced the in Vitro Diagnostic Devices Regulation 2017/746 (IVDR) and forever changed the IVD regulatory framework. Since then, IVD manufacturers, notified bodies (NBs), distributors, and Competent Authorities have been scrambling to make sense of the new requirements and scale their operations to achieve and maintain compliance under the new regulations. Final transition deadlines are approaching and IVD companies need to take action.
EU IVDR Classification
Most IVDs were reclassified under the IVDR, which means you will likely face more rigorous conformity assessment requirements. Before you can identify those requirements, you must accurately classify your device according to IVDR implementing and classification rules. We will help you refine your device’s intended purpose, if needed, and assess your device’s classification according to the IVDR.
EU CE Marking for IVDs
New IVDs require the applicable IVDR conformity before market placement. Eligible legacy devices may use Article 110 transitions, while devices already lawfully placed on the market have separate continued-availability rules. When your IVD requires IVDR CE Marking will depend on your device’s legacy status and device classification. To ensure compliance by your IVD’s deadline, Pure Global can perform a thorough evaluation of your technical documentation and clinical evidence to assess your IVDR readiness and develop a regulatory strategy for IVDR CE Marking.
EU IVDR PER Performance Evaluation Report
The Performance Evaluation Report (PER) is an essential part of your Technical Documentation for CE Marking that must be updated continuously throughout the product life cycle. It includes individual reports for scientific validity, analytical performance, and clinical performance data, as well as explanations of the methods you used to gather clinical evidence, perform literature searches, etc. We can help you plan your performance evaluation process and collaborate with your team to maintain compliance with performance evaluation requirements.
EU PMPF Post-Market Performance Follow-Up for IVDs
Post-Market Performance Follow-up (PMPF) is a new but critical requirement introduced under the IVDR. PMPF is an ongoing process to collect and evaluate performance and scientific data from the use of your device on the market. PMPF data supports your Performance Evaluation (PE), risk management, and Post-Market Surveillance (PMS) activities. We can help you design a PMPF plan and execute PMPF activities, including PMPF studies, throughout the device life cycle.
EU Eudamed Medical Device & IVD Registration
Mandatory use of the first four EUDAMED modules began on 28 May 2026. Manufacturers and other applicable economic operators must maintain actor and device data, including Unique Device Identifier (UDI) information, under the current transition rules. We can help obtain UDI codes, prepare records, and build EUDAMED maintenance into the regulatory process.
EU MDR and IVDR Post-Market Surveillance (PMS)
Post-Market Surveillance requirements expanded under the EU MDR and EU IVDR. Our team of regulatory professionals specializes in helping medical device manufacturers develop and implement compliant, efficient, and risk-based PMS strategies tailored to their products. We provide hands-on guidance to ensure that your PMS plans, reports, and procedures meet the stringent EU MDR and IVDR requirements.
EU Representative European Authorized Representative for Medical Devices & IVD Companies
If you’re a non-EU manufacturer of medical devices or in-vitro diagnostics (IVDs) looking to sell in Europe, you need a trusted EU Authorized Representative (EU AR) to ensure compliance with the EU MDR and EU IVDR. The EU AR performs its Article 11 verification and cooperation tasks under a written mandate; the manufacturer retains its own evidence, vigilance and postmarket obligations. With offices throughout Europe staffed by experienced medical device and IVD regulatory professionals, we can act as your regulatory and commercial partner.
CE Mark Notified Body Search and Selection
Finding the right Notified Body can be challenging due to limited availability, long review timelines, and varying expertise across different Notified Bodies. Choosing the wrong Notified Body can result in delays, increased costs, or even non-approval of your product, significantly impacting your market access. With our expert support, you can efficiently identify, evaluate, and engage with the best Notified Body for your device, ensuring a smooth certification process and faster time-to-market.
EU MDR & IVDs PRRC Person Responsible for Regulatory Compliance
Under the EU MDR and IVDR, medical device manufacturers selling in Europe must appoint a Person Responsible for Regulatory Compliance (PRRC) who is responsible for overseeing regulatory compliance, quality management, and post-market surveillance. The PRRC must have a degree in a relevant discipline plus one year of regulatory experience or at least four years of experience in regulatory affairs or quality management for medical devices.
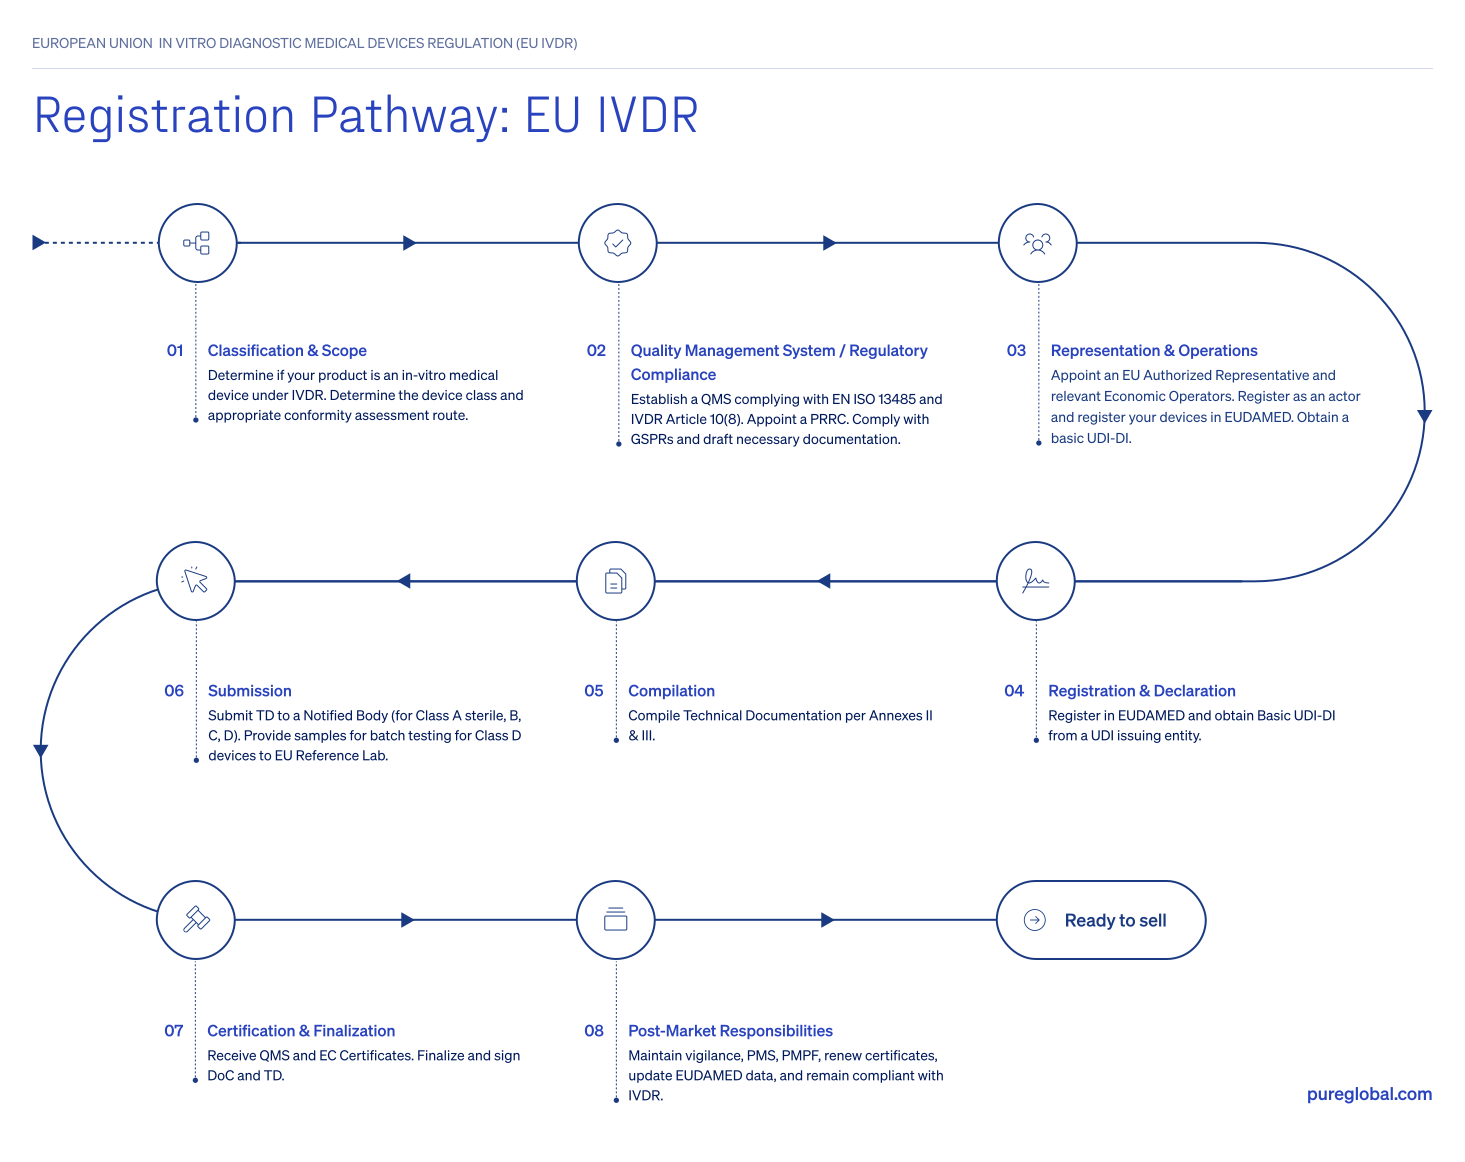
IVDR legacy transition milestones
| IVDR class | Latest application milestone for the relevant extended route | Written agreement milestone | End of eligible placement period |
|---|---|---|---|
| D | May 26, 2025 | September 26, 2025 | December 31, 2027 |
| C | May 26, 2026 | September 26, 2026 | December 31, 2028 |
| B and A sterile | May 26, 2027 | September 26, 2027 | December 31, 2029 |
These are conditional transitions, not automatic extensions for every IVD. The QMS milestone was May 26, 2025, and additional conditions apply. As of this review, the Class D and C application/agreement milestones have passed: confirm documentary eligibility rather than proposing to meet them later. Use the Commission transition guidance and actual Article 110 conditions.
Build a product-level evidence plan connecting GSPR mapping methodology to the correct IVDR Annex I, performance evaluation, risk management, labels and PMS reporting. IVDR-specific scientific validity, analytical and clinical performance must be supported. Follow the current Commission guidance and IVDR legal framework.
For regulatory terminology and the authority framework, see the EU MDR.
 Regulatory Update · Oct 11, 2026EU First Device JCA SelectionRegulatory Update · Sep 14, 2026COMBINE: Combined-Study Safety ReportingRegulatory Update · Sep 14, 2026MDCG Rule 7: IVD Quality ControlsRegulatory Update · Aug 26, 2026EU AI Act Annex I Date 2028Case Study · Aug 11, 2026Basic UDI-DI Grouping for Contrast DevicesCase Study · Aug 11, 2026Class III Dermal Fillers Under the EU MDRCase Study · Aug 5, 2026EU Authorized Representative for Surgical RobotsBlog · Oct 19, 2026MDR submission completenessBlog · Oct 14, 2026SaMD vs SiMD GuideBlog · Oct 12, 2026EUDAMED registration clocks
Frequently asked questions
A scoped IVDR project can cover IVD qualification and classification, gap assessment, quality-system planning, technical documentation, performance evaluation, labeling, post-market performance follow-up, EUDAMED and UDI readiness, EU Authorized Representative coordination, and Notified Body submission support. The exact work depends on the device class and current evidence.
Class B, C, and D IVDs generally require Notified Body conformity assessment. Class A devices normally self-declare, although sterile Class A devices need Notified Body involvement for the aspects related to establishing, securing, and maintaining sterile conditions. Classification and applicable conformity-assessment annexes should be confirmed before engaging a body.
Start with a controlled intended-purpose statement, IVDR classification rationale, conformity-assessment route, ISO 13485 gap analysis, performance-evaluation plan, analytical and clinical performance evidence, risk management, labeling, PMS and PMPF plans, PRRC and EU representative arrangements, and an EUDAMED and UDI plan. Resolve major evidence gaps before reserving review capacity.
Single Process,
Multiple Markets
When you partner with Pure Global, a single registration process opens doors to multiple countries. Our global subsidiaries make this streamlined path possible.








Let's Talk,
Anywhere You Are.
Whether looking for more information or ready to partner with us, we're here to guide you through every step of the regulatory process.
Contact us